What Certifications Are Needed for Medical Beds in 2026?

Time:2026-09-30 Author:Aria
0%

Choosing a medical bed is not only a matter of comfort, dimensions, or price. Buyers also need evidence that its design, electrical systems, and manufacturing controls suit the intended use. The question “what certifications are needed for medical beds” has no single answer for every product or market. Requirements can vary with bed features, patient population, sales region, and whether the bed is classified as a medical device. Details matter.

Several frameworks may be relevant in 2026. IEC 60601-2-52 addresses basic safety and essential performance for adult medical beds, while IEC 60601-1 covers general medical electrical equipment safety. ISO 13485 concerns quality management systems, and ISO 14971 provides a framework for managing medical-device risks. These standards are not interchangeable certificates. CE marking and U.S. FDA requirements also follow different regulatory pathways; FDA clearance, for example, should not be casually described as a certification. Small wording differences can mislead a purchasing team.

A useful review asks what applies to this exact bed, in this exact market. Request current test reports, declarations, device registrations or clearances where applicable, and documentation linking them to the model number. Check the powered adjustments, side rails, control handset, and labeling—not just the brochure. Ask who issued each document and whether its scope covers the bed configuration being offered. A logo alone proves little. Even a tidy compliance folder can leave unanswered questions, so independent review may be worthwhile. The sections below explain the main standards, regional differences, and practical checks buyers can use before making a decision.
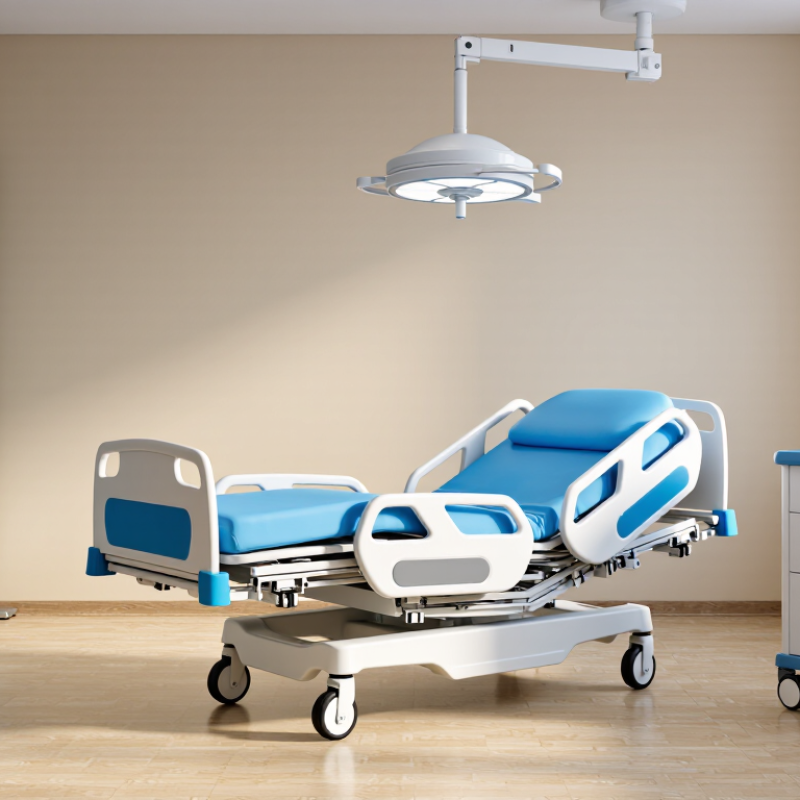
What Certifications Are Needed for Medical Beds in 2026?

Determine the Medical Bed’s Classification and Target Markets

What Certifications Are Needed for Medical Beds in 2026?

Determine the Medical Bed’s Classification and Target Markets

A medical bed’s certification path starts with classification, not paperwork. Define its intended use, users, patient risks, and operating environment. A basic manually adjustable bed may face different controls from a powered bed with integrated weighing, fall detection, or positioning software. Claims change everything. Classification can also vary by country, even when the product design remains unchanged.

For the United States, the manufacturer may need FDA establishment registration, device listing, and possibly premarket clearance. The exact route depends on the device’s intended use and substantial equivalence. In the European Union, compliance with the Medical Device Regulation may require CE marking, technical documentation, clinical evaluation, and notified-body involvement. Other markets, including the United Kingdom, Canada, and Australia, apply separate registration systems and transition rules. Do not treat CE, FDA, or ISO documents as interchangeable.

Testing should match the final design. Common evidence may include IEC 60601-1 electrical safety, IEC 60601-1-2 electromagnetic compatibility, and IEC 60601-2-52 for medical beds. ISO 14971 supports risk management, while ISO 13485 strengthens the quality system. These standards do not automatically grant market approval. A test report is not a license. That distinction is easy to miss.

I would confirm classification with local regulatory specialists before changing labels, software, or accessories. Small changes can create new obligations. In practice, the target market should be selected before certification testing begins, although real projects often do this too late.

Identify Required Regulatory Registrations and Market Approvals

Medical-bed requirements in 2026 depend on the destination market, intended use, and product design. A standard ward bed may follow different pathways from a bed with powered positioning or specialized safety functions. In the United States, manufacturers may need FDA establishment registration and device listing, plus a market authorization pathway such as 510(k), De Novo, or PMA when applicable. Registration alone does not mean the device is FDA-approved. That distinction matters.

For the European Union, a medical bed placed on the market generally needs to meet the Medical Device Regulation, carry CE marking, and involve a notified body when its classification requires one. Other markets may require local registration, an in-country representative, importer documentation, or language-specific labels. Technical evidence commonly includes risk-management records, electrical-safety and electromagnetic-compatibility testing, and relevant bed-safety testing. Requirements can change, so verify the current rules with the regulator or a qualified compliance professional.

Tips: Map each target country before production. Keep intended-use statements, test reports, labels, and registration records consistent. Check every accessory too. A small change, such as adding a control feature, may affect classification or testing. It is easy to overlook that detail, even with a strong compliance plan.

Review Applicable Quality Management System Requirements

What Certifications Are Needed for Medical Beds in 2026?

In 2026, medical bed manufacturers need more than a product certificate. Their quality management system must control design, production, testing, and post-market feedback. ISO 13485:2016 remains the main international benchmark for medical device quality systems. However, certification alone does not guarantee market access. Requirements still depend on the destination country, device classification, and bed configuration.

In the United States, manufacturers should prepare for the Quality Management System Regulation, which aligns more closely with ISO 13485. A strong system should include document control, supplier evaluation, employee training, complaint handling, and corrective actions. Keep clear records. Auditors may examine a single design change, supplier issue, or service report in detail. For powered beds, electrical safety and performance standards may also apply, but these are not substitutes for a quality management system.

Risk Management and Practical Readiness

Risk management deserves practical attention. ISO 14971 can support the identification of hazards such as unintended movement, patient entrapment, unstable accessories, and control failure. It is generally a process standard, not a standalone product certification. Manufacturers should connect risk files with testing, instructions, maintenance procedures, and field data. This connection is often weaker than expected. A polished manual cannot repair incomplete training records. Regular internal audits, realistic mock inspections, and honest review of near misses can reveal gaps before regulators or customers do.

Verify Electrical, Mechanical, and Usability Safety Standards

What Certifications Are Needed for Medical Beds in 2026?

Verify Electrical, Mechanical, and Usability Safety Standards

Medical bed certification in 2026 depends on the destination market and bed classification. However, electrical safety usually starts with IEC 60601-1. Testing should also address electromagnetic compatibility under IEC 60601-1-2. These checks examine leakage current, insulation, grounding, and interference. A qualified laboratory should review the complete bed, including motors, control boxes, handsets, and charging systems.

Mechanical safety needs equal attention. IEC 60601-2-52 covers important requirements for medical beds and their accessories. Assessors commonly examine load capacity, stability, side-rail strength, mattress-platform movement, and entrapment risks. Repeated movement tests matter. A bed may pass a static load test but fail after months of daily adjustment. That detail is easy to overlook.

Usability certification is not just a user-interface exercise. IEC 62366-1 supports a process for identifying use-related risks, especially during transfers, emergency positioning, and control selection. ISO 14971 can help connect these risks with the wider safety file. Manufacturers should maintain evidence through design records, risk analysis, test reports, and production controls under ISO 13485. Local registration rules may add electrical, mechanical, labeling, or clinical documentation. Requirements can change. A checklist may look complete and still hide a gap. Independent review remains worthwhile before market release.

Maintain Technical Documentation, Labeling, and Post-Market Compliance

What Certifications Are Needed for Medical Beds in 2026?

For medical beds, certification is only the starting point. Manufacturers should maintain technical documentation throughout the product’s life. IEC 60601-1 addresses basic electrical safety. IEC 60601-2-52 focuses on the safety and performance of medical beds. ISO 14971 supports risk management, while ISO 13485 controls the quality system behind production and servicing.

Documentation must match the actual bed. Keep drawings, software versions, test reports, risk files, usability evidence, and supplier records current. A changed actuator or side-rail design may require a fresh risk review. Do not rely on an old certificate. That approach can fail during an audit. In the United States, FDA’s 2023 medical device reporting data recorded more than two million adverse-event reports across devices. The figure shows why traceable records matter, even when a bed appears mechanically simple.

Labeling also needs practical attention. Include load limits, lock positions, electrical warnings, cleaning instructions, and safe-use symbols. Labels should remain readable after repeated disinfection. Post-market compliance requires complaint trending, incident assessment, corrective actions, and periodic safety reviews. The World Health Organization’s Global Patient Safety Action Plan notes that about one in ten patients experiences harm in healthcare, with more than half considered preventable. A weak feedback process can hide recurring entrapment, brake, or control-panel problems. Documentation is often treated as paperwork. It should instead tell the product’s real safety story. Even experienced teams occasionally miss small changes. That is worth reviewing.

What Certifications Are Needed for Medical Beds in 2026? — Maintain Technical Documentation, Labeling, and Post-Market Compliance

Key point: There is no single worldwide “medical bed certificate.” Applicable approvals depend on the intended purpose, design, powered functions, risk classification, and markets where the bed is placed on the market.

Compliance area Market or reference Typical evidence or action 2026 status and practical consideration
Device classification and market authorization All markets Document the intended purpose, intended users and settings, bed functions, accessories, and rationale for the device classification. Identify each applicable market’s conformity-assessment route. Market-specific
Classification cannot be determined from the product name alone. Powered features or medical claims may affect the applicable route.
EU conformity assessment and CE marking European Union — Regulation (EU) 2017/745 (MDR) Maintain technical documentation, risk-management records, clinical evaluation as applicable, a post-market surveillance plan, and an EU declaration of conformity. Apply CE marking after the applicable conformity-assessment requirements are met. Required for applicable devices
The route depends on MDR classification. A notified body is required when the applicable class or device features require one; some Class I devices may use manufacturer self-declaration.
Electrical safety IEC 60601-1 and relevant national or regional adoptions For electrically powered medical beds, maintain test evidence and a safety assessment covering hazards such as electric shock, mechanical hazards, temperature, and essential performance where applicable. Applicability depends on design
Use the edition and national adoption applicable to the target market and conformity route. A test report is not, by itself, market authorization.
Electromagnetic compatibility IEC 60601-1-2 and applicable regional requirements Keep electromagnetic emissions and immunity test evidence for applicable powered equipment, including relevant configurations, accessories, and operating modes. Applicable to relevant electrical devices
Assess whether electromagnetic disturbances could affect movement, controls, alarms, or other safety-related functions.
Particular requirements for medical beds IEC 60601-2-52 and applicable national or regional adoptions Use the applicable particular standard as part of the safety assessment for medical beds within its scope. Retain evidence addressing bed-specific hazards and performance requirements. Scope-dependent
Confirm the product falls within the standard’s scope and check the current edition and its recognition or adoption in each target market.
Quality management system ISO 13485:2016; jurisdictional requirements Maintain controlled procedures and records for design and development, supplier controls, production, traceability, complaint handling, corrective action, and regulatory reporting as applicable. Regulatory or voluntary, depending on market
ISO 13485 certification is not a universal product approval. A regulator or conformity-assessment route may require a compliant QMS or specific evidence.
Risk management ISO 14971:2019; applicable regulatory requirements Keep a lifecycle risk-management file covering hazard identification, risk evaluation, risk controls, verification, residual-risk evaluation, and production and post-production information. Core technical documentation
Consider foreseeable misuse and hazards such as entrapment, falls, unintended movement, instability, and failures of controls or power systems.
U.S. device status and establishment obligations United States — FDA; 21 CFR 880.5100 Confirm the exact device type, classification, and any applicable premarket requirements. Complete FDA establishment registration and device listing when required, and retain records supporting the regulatory determination. Verify the specific device
FDA lists hospital beds under 21 CFR 880.5100 as Class I. Any 510(k) exemption is subject to regulatory limitations and should be verified for the exact device and intended use.
U.S. quality system requirements United States — FDA Quality Management System Regulation (QMSR), 21 CFR Part 820 Assess the manufacturer’s applicable quality-system obligations and maintain required quality records and procedures. Effective February 2, 2026
The QMSR incorporates ISO 13485:2016 by reference. Check the regulation for its scope, applicable exemptions, and requirements for the specific organization and device.
Labeling and instructions for use All target markets; examples include EU MDR Annex I and U.S. FDA labeling requirements Provide the required manufacturer and device identification, intended purpose, warnings, precautions, operating instructions, maintenance information, and other market-required particulars. Use required languages for each market. Market-specific and safety-critical
Ensure the information matches the cleared or declared intended use and the actual configuration. Keep label and instruction revisions controlled.
Unique device identification and traceability Markets with UDI requirements, including the EU and United States Determine whether the device and its packaging require a UDI. Maintain applicable identifier, labeling, database-submission, and traceability records. Applicability and deadlines vary
Check device-class rules, exemptions, packaging levels, and market-specific database obligations rather than assuming one global UDI rule.
Post-market surveillance and vigilance EU MDR, U.S. FDA, and other market rules Maintain complaint handling, incident assessment, trend monitoring, investigation, corrective and preventive action, and required safety-reporting processes. Prepare applicable periodic or summary reports. Ongoing obligation
Reporting timelines and report types differ by jurisdiction. Keep evidence that complaints and field information are reviewed and acted upon.
Changes, field actions, and record retention All target markets Control design, supplier, software, manufacturing, and labeling changes. Assess whether a change affects conformity, classification, authorization, or existing registrations. Document field corrections and safety actions. Lifecycle control
Retain technical, manufacturing, distribution, and post-market records for the periods required by the applicable regulations.
Other national registration or listing Examples: United Kingdom, Canada, Australia Check local registration, licensing, conformity-assessment, importer, sponsor, or device-listing obligations before placing the product on the market. Separate national checks required
For example, UK requirements are administered under the UK medical-device framework; Canada and Australia have their own licensing or inclusion rules. Requirements depend on device class and supply-chain role.

Planning note: Standards and regulatory rules may be revised or adopted differently by jurisdiction. Before market launch, verify current editions, recognized or harmonized status, classification, exemptions, labeling rules, and transition dates with the relevant regulator or conformity-assessment body.

FAQS

What determines a medical bed’s certification pathway in 2026?

Classification comes first. Consider intended use, users, patient risks, software, accessories, and the operating environment.

Does one certification cover every country?

No. Requirements differ by market. A document accepted in one country may not support approval elsewhere.

What may be required for sales in the United States?

Manufacturers may need establishment registration, device listing, and market authorization. The route depends on intended use and product similarity.

What is generally needed for the European market?

The bed may need compliance with the regional medical-device framework, technical documentation, clinical evaluation, and independent assessment.

Which safety tests commonly apply to powered medical beds?

Electrical safety, electromagnetic compatibility, and medical-bed safety tests are commonly expected. Test the complete bed, including motors and hand controls.

What mechanical risks should testing examine?

Review load capacity, stability, side-rail strength, platform movement, and patient entrapment risks. Repeated movement tests matter.

Why is usability assessment important?

Users may select the wrong control during transfers or emergencies. Usability analysis should identify confusing buttons, poor labels, and risky actions.

Can adding software or an accessory change certification requirements?

Yes. A weighing function, fall alert, or new handset may change classification, testing, or documentation duties.

Is a test report the same as market approval?

No. A test report provides evidence, not permission to sell. That distinction is easy to miss.

How can manufacturers reduce certification problems?

Select target markets before testing. Keep labels, risk files, reports, and registration records consistent. A checklist can still hide a gap.

Conclusion

Determining what certifications are needed for medical beds in 2026 begins with identifying the bed’s classification, intended use, and target markets. Requirements may differ depending on whether the product is intended for hospitals, long-term care facilities, home care, or specialized clinical environments. Manufacturers should identify the appropriate regulatory registrations, conformity assessments, and market approvals before commercial distribution.

A complete compliance plan should also address quality management system requirements, electrical and mechanical safety, electromagnetic compatibility, stability, durability, and usability. Technical files should include risk management records, testing evidence, clinical or performance support where applicable, labeling, instructions for use, and traceability information. After launch, companies must maintain complaint handling, incident reporting, corrective action, and periodic product monitoring processes. By coordinating classification, approvals, quality controls, safety verification, documentation, and post-market activities, manufacturers can support safe medical bed performance and maintain access to their intended markets in 2026.
Aria

Aria

Aria is a dedicated marketing professional with a deep passion for innovative strategies and a keen understanding of our company's product offerings. With a wealth of experience in the industry, Aria excels at crafting engaging content that highlights the unique features and benefits of our......